FDA Publishes Draft Guidance on Essential Drug Delivery Outputs for Drug Delivery Devices
FDA has published draft guidance on Essential Drug Delivery Outputs (EDDOs) for drug delivery devices and combination products. Learn how the recommendations may impact design controls, verification, validation, and change management activities.
MedTech Europe Calls for Clarification on EDPB’s Proposed DPIA Template
MedTech Europe has responded to the EDPB consultation on the proposed DPIA template, calling for greater flexibility and warning against additional compliance burdens for medical device and digital health manufacturers.
Swissmedic Publishes swissdamed Business Rules Version 4.0
Swissmedic has published swissdamed Business Rules Version 4.0, introducing new Master UDI-DI requirements, UDI registration rules, and data management obligations for medical device manufacturers in Switzerland.
FDA Updates Guidance on Manufacturer Communications with Payors: What Medical Device Companies Need to Know
The FDA has updated its draft guidance on medical device manufacturer communications with payors, health economic information, and investigational products.
FDA Updates eSTAR Program: Human Factors Content Added and Expanded PreSTAR Functionality
The FDA has released eSTAR Version 7.0, integrating new Human Factors content and expanding PreSTAR capabilities. Learn how these changes may affect medical device manufacturers preparing FDA submissions.
WHO Revises International Biological Reference Standards Guidance: Key Considerations for IVD and Biological Product Manufacturers
The WHO has adopted revised recommendations for International Biological Reference Standards (WHO IBRS), introducing updated guidance on commutability, calibration, secondary standards and International Units. Learn what IVD and biological product manufacturers need to know.
MedTech Europe Publishes IVD-Specific Recommendations for the MDR/IVDR Revision
MedTech Europe has published new recommendations for the MDR/IVDR revision, calling for a more proportionate regulatory framework, support for innovation, and improved access to in vitro diagnostics across Europe.
ICH Adopts E6(R3) Annex 2: New Global Guidance for Decentralised Trials, Digital Technologies and Real-World Data
ICH has adopted E6(R3) Annex 2, introducing new Good Clinical Practice guidance for decentralised clinical trials, digital health technologies and real-world data. Learn what it means for medical device manufacturers.
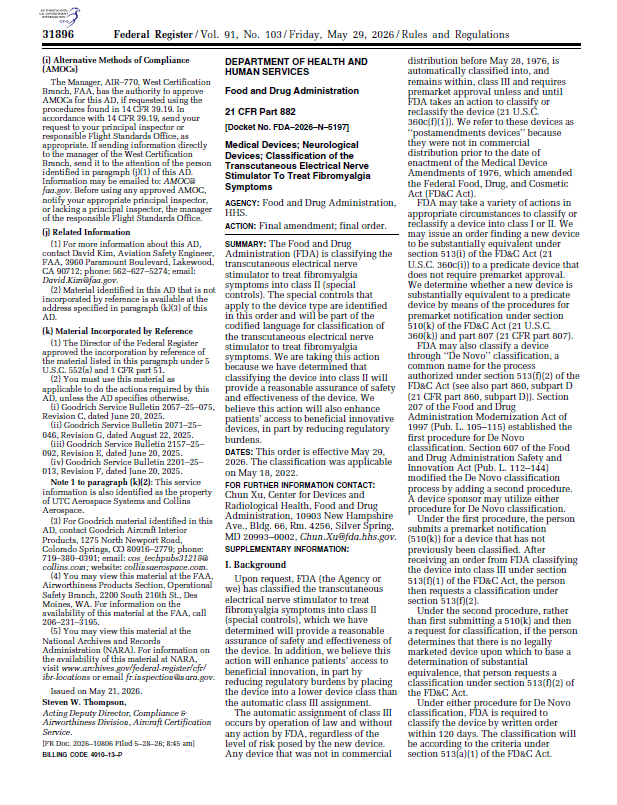
FDA Establishes Class II Classification for TENS Devices Intended to Reduce Fibromyalgia Symptoms
FDA has established a new Class II classification with special controls for transcutaneous electrical nerve stimulators (TENS) intended to treat fibromyalgia symptoms, creating a new regulatory pathway for manufacturers.
Team-NB Publishes Position Paper on the Transfer of IVDR Applications and Legacy Device Surveillance Between Notified Bodies
Team-NB has published a position paper introducing a template agreement for transferring IVDR applications and legacy device surveillance between notified bodies.
Team-NB Publishes Updated Consensus Document on the MDR Certification Process
Team-NB has published Version 2 of its MDR Certification Process Consensus Document, providing detailed guidance on pre-application, application, conformity assessment and surveillance activities for medical device manufacturers under EU MDR 2017/745.
FDA Reports 1,284 Devices Granted Breakthrough Device Designation as of March 2026
The FDA has reported 1,284 Breakthrough Device designations and 198 market-authorised Breakthrough Devices as of March 2026. Learn what this means for medical device manufacturers and U.S. market access strategies.
FDA Adds Five Additional Unclassified Medical Devices to 510(k) Enforcement Discretion Policy
FDA has updated its guidance on unclassified medical devices, adding five additional device types to its 510(k) enforcement discretion policy. Learn what this means for manufacturers and ongoing FDA compliance obligations.
FDA Adopts ICH M11 CeSHarP Guidance: What Sponsors Need to Know About the Future of Clinical Trial Protocols
The FDA has adopted ICH M11 CeSHarP, introducing a harmonized framework for clinical trial protocols. Discover the impact on sponsors and global studies.
European Commission Publishes Third Version of Living Guidelines on the Responsible Use of Generative AI in Research
The European Commission has released Version 3 of its Living Guidelines on the Responsible Use of Generative AI in Research. Learn what the new recommendations mean for medical device manufacturers, clinical evaluation, data protection and research activities.
MedTech Europe Responds to Consultation on the Revision of the EU Cybersecurity Act
MedTech Europe responds to the EU Cybersecurity Act consultation, highlighting key implications for medical device and IVD manufacturers, including certification, vulnerability management, and supply chain security.
FDA Finalizes Risk-Based Guidance on Human Factors Information in Medical Device Submissions
The FDA has published its final guidance on Human Factors information in medical device submissions, introducing a new risk-based framework and three Human Factors Submission Categories. Learn what manufacturers need to know.
FDA Replaces 25-Year-Old Bioequivalence Statistical Guidance with Major Update
The FDA has released a major update to its Bioequivalence Statistical Guidance, replacing the 2001 version. The new document introduces adaptive designs, modern statistical methods, guidance for highly variable drugs, NTI products, missing data handling and model-based approaches for generic drug development.
Australia Updates Guidance on UDI Compliance Timeframes for Medical Devices
The TGA has updated its guidance on UDI compliance timeframes for medical devices in Australia, including clarifications on legacy devices, EU MDR transitions, consignment stock, and Surgical Loan Kits.
Proposed MDR Article 50 Raises Concerns Over Notified Body Independence and Financial Sustainability
A new legal opinion analyses the European Commission’s proposed MDR Article 50, highlighting potential impacts on notified bodies, SMEs, certification costs, and conformity assessment access.