
ANVISA Launches Public Consultation to Review the Regulatory Framework for Medical Device Vigilance
ANVISA has launched a public call for contributions to support the revision of Brazil's medical device vigilance regulatory framework, seeking feedback from medical device registration holders on post-market surveillance requirements and future regulatory developments.

TGA Introduces Australia's UDI System with Phased Implementation Through 2029
The TGA has introduced Australia's Unique Device Identification (UDI) system with a phased implementation through 2029. Learn about the implementation timeline, AusUDID requirements, and what medical device and IVD manufacturers need to prepare.

FDA Classifies Endoscopic Traction Devices as Class II Medical Devices
FDA classifies the Endoscopic Traction Device as a Class II medical device with Special Controls, setting performance, safety, and labeling requirements applicable to medical device and IVD manufacturers.

MDCG publishes guidance on the transition from the ‘EC REP’ to the ‘EU REP’ symbol
The MDCG has published an appendix to MDCG 2021-5 clarifying the transition from the 'EC REP' to the 'EU REP'symbol under EN ISO 15223-1, including a five-year transition period for medical device and IVD manufacturers.
Swissmedic publishes updated swissdamed Playground User Guide
Swissmedic has published Version 3.0 of the swissdamed User Guide – Playground, providing updated instructions for testing actor registration and UDI management before using the production environment.

European Parliament briefing highlights implementation of the EU Artificial Intelligence Act
A new European Parliament briefing provides an overview of the EU Artificial Intelligence Act, including its risk-based framework, implementation timeline, and key provisions relevant to AI systems, including those used in medical devices.
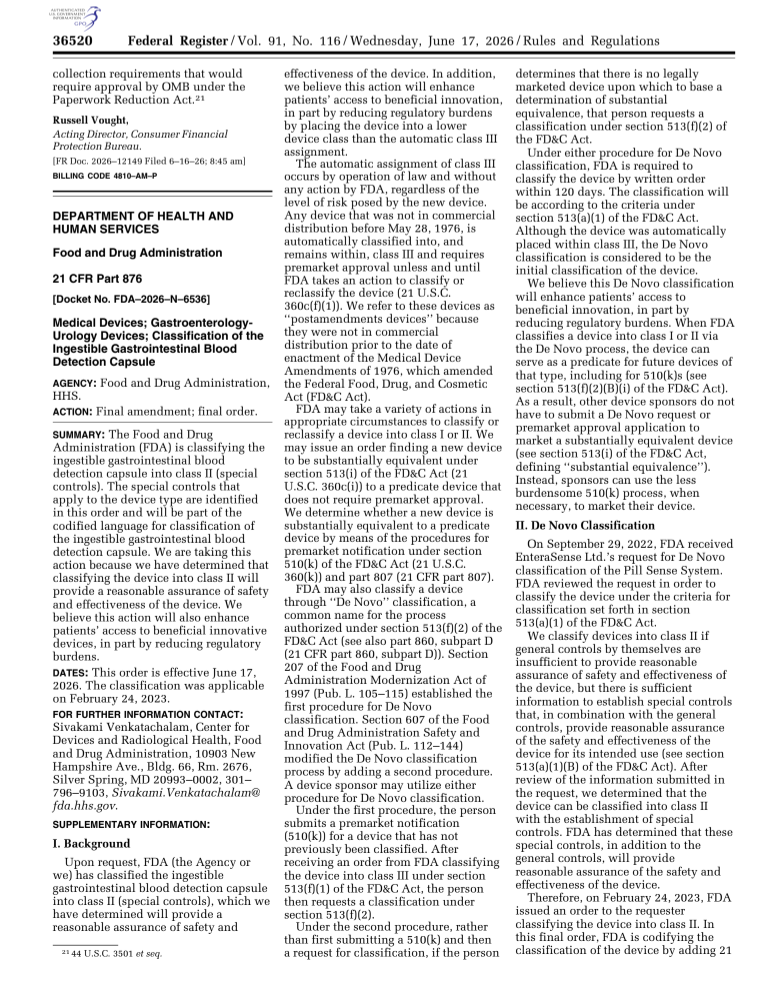
FDA Classifies Ingestible Gastrointestinal Blood Detection Capsule as a Class II Medical Device
The FDA has classified the ingestible gastrointestinal blood detection capsule as a Class II medical device, establishing special controls to support the safety and effectiveness of this innovative diagnostic technology.

FDA Classifies Radiological Machine Learning-Based Quantitative Imaging Software with Predetermined Change Control Plans as Class II Devices
The FDA has classified radiological machine learning-based quantitative imaging software with predetermined change control plans as Class II medical devices, establishing special controls to support the safe development and regulatory oversight of AI-enabled imaging software.
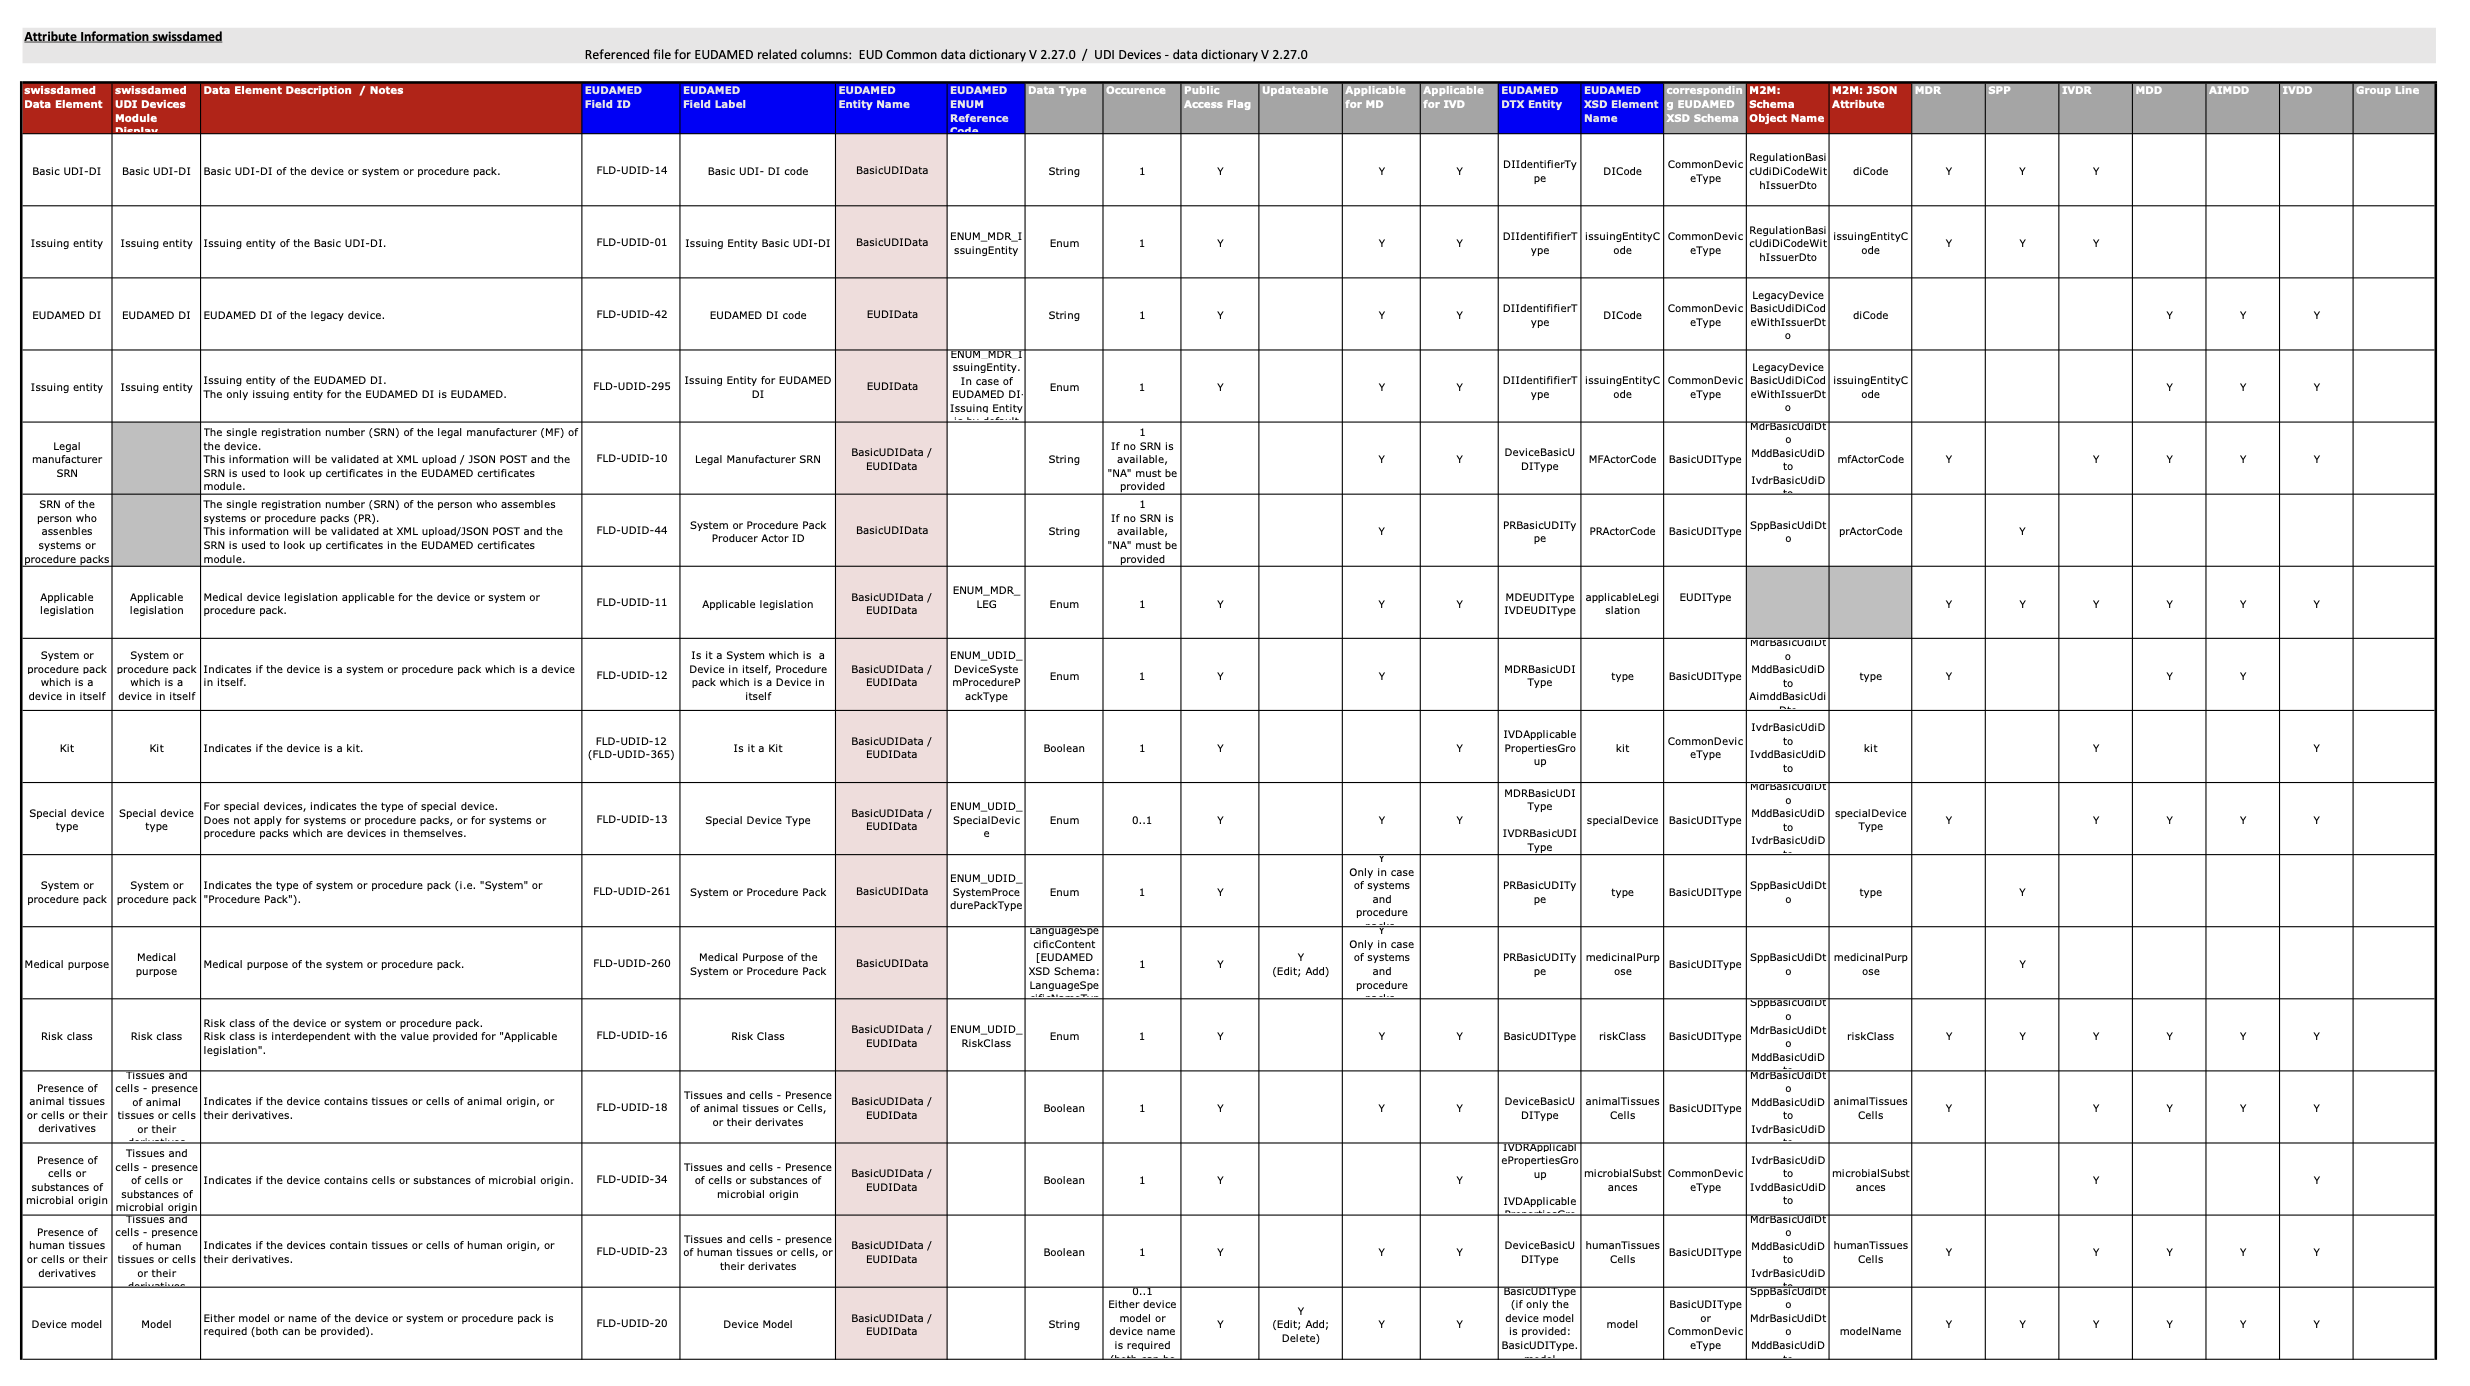
Swissmedic Publishes Version 2.0 of the swissdamed UDI Data Dictionary
Swissmedic has published Version 2.0 of the swissdamed UDI Data Dictionary, introducing new data elements and clarifying requirements for the registration and management of UDI information for medical devices in Switzerland.
Swissmedic Publishes Guidance on the Procurement of Medical Devices in Healthcare Institutions
Swissmedic has published Version 6.0 of its Information Sheet – Procurement of Medical Devices in Healthcare Institutions, providing guidance on the procurement of medical devices and IVDs in Switzerland, including conformity documentation, certificate verification and the use of swissdamed.

Swissmedic Publishes User Guide for the swissdamed UDI Devices Module
Swissmedic has published Version 3.0 of the swissdamed User Guide – UDI Devices Module, introducing new functionalities for device registration and management, including Machine-to-Machine (M2M) processes, manual UDI-DI management and Master UDI-DI requirements.

MDCG Publishes Position Paper on the Management of SS(C)Ps in EUDAMED After Mandatory Use
The MDCG has published MDCG 2026-4, providing clarification on the management of Summary of Safety and Clinical Performance (SSCP) and Summary of Safety and Performance (SSP) documents in EUDAMED following the mandatory use of the platform’s first modules.

MedTech Europe Supports MDR and IVDR Simplification Proposal Focused on More Proportionate Lifecycle Oversight
MedTech Europe has published its position on the European Commission’s proposal to simplify the MDR and IVDR, supporting a more proportionate and risk-based approach to oversight, surveillance and assessment activities throughout the lifecycle of medical devices.
Swissmedic Updates Formal Requirements Guidance for Medicinal Product Authorisation Applications
Swissmedic has published Version 21.0 of its Formal Requirements guidance document, introducing updates and clarifications relating to medicinal product authorisation applications, documentation requirements, biosimilars, Article 13 TPA submissions and combination products incorporating medical devices.
FDA Publishes Draft Guidance on Forms FDA 3542a and FDA 3542 for Orange Book Patent Submissions
FDA has published draft guidance on Forms FDA 3542a and FDA 3542, clarifying requirements for Orange Book patent listings, electronic patent submissions, and the management of patent information throughout the medicinal product lifecycle.

ENISA Publishes 2026 Report on the State of SBOM Adoption Across Europe
ENISA has published its SBOM Adoption State of Play – 2026 report, providing insights into how organisations across Europe are implementing Software Bills of Materials (SBOMs) in response to the Cyber Resilience Act (CRA) and identifying key challenges affecting adoption.

CIOMS Publishes Updated Glossary of ICH Terms and Definitions
The Council for International Organizations of Medical Sciences (CIOMS) has published Version 10 of its Glossary of ICH Terms and Definitions, consolidating terminology from current ICH guidelines and supporting greater consistency in regulatory communication across the life sciences sector.
MHRA Publishes AI Airlock Phase 2 Report on Regulatory Challenges for AI Medical Devices
The MHRA has published the Phase 2 report of its AI Airlock programme, highlighting regulatory challenges related to AI medical devices, AI-powered IVDs, performance evaluation, post-market surveillance and lifecycle change management.
EMA Updates Guidance on Electronic Submission of Medicinal Product Data under PMS and ISO IDMP
The EMA has published version 3.6 of its guidance on the implementation of ISO IDMP standards through the Product Management Services (PMS), introducing updates related to data submission, data enrichment, and requirements applicable to marketing authorisation holders.
Health Canada summarizes feedback on Draft Guidance for Decentralized Clinical Trials
Health Canada published feedback from its public consultation on Decentralized Clinical Trials, including topics related to digital technologies and data management.